methylseg 0.1.8__tar.gz
This diff represents the content of publicly available package versions that have been released to one of the supported registries. The information contained in this diff is provided for informational purposes only and reflects changes between package versions as they appear in their respective public registries.
- methylseg-0.1.8/LICENSE +28 -0
- methylseg-0.1.8/PKG-INFO +320 -0
- methylseg-0.1.8/README.md +282 -0
- methylseg-0.1.8/methylseg/__init__.py +38 -0
- methylseg-0.1.8/methylseg/cli.py +32 -0
- methylseg-0.1.8/methylseg/data_manager.py +84 -0
- methylseg-0.1.8/methylseg/helper_classes.py +816 -0
- methylseg-0.1.8/methylseg/methyl_segmentor.py +510 -0
- methylseg-0.1.8/methylseg/methyl_state_analyzer.py +769 -0
- methylseg-0.1.8/methylseg/methyl_state_assigner.py +2643 -0
- methylseg-0.1.8/methylseg/methylseg_config.py +1088 -0
- methylseg-0.1.8/methylseg/methylseg_hmm.py +455 -0
- methylseg-0.1.8/methylseg/methylseg_pathway.py +1965 -0
- methylseg-0.1.8/methylseg/utils.py +1125 -0
- methylseg-0.1.8/methylseg.egg-info/PKG-INFO +320 -0
- methylseg-0.1.8/methylseg.egg-info/SOURCES.txt +20 -0
- methylseg-0.1.8/methylseg.egg-info/dependency_links.txt +1 -0
- methylseg-0.1.8/methylseg.egg-info/entry_points.txt +2 -0
- methylseg-0.1.8/methylseg.egg-info/requires.txt +29 -0
- methylseg-0.1.8/methylseg.egg-info/top_level.txt +1 -0
- methylseg-0.1.8/pyproject.toml +59 -0
- methylseg-0.1.8/setup.cfg +4 -0
methylseg-0.1.8/LICENSE
ADDED
|
@@ -0,0 +1,28 @@
|
|
|
1
|
+
BSD 3-Clause License
|
|
2
|
+
|
|
3
|
+
Copyright (c) 2026, Clement Lab
|
|
4
|
+
|
|
5
|
+
Redistribution and use in source and binary forms, with or without
|
|
6
|
+
modification, are permitted provided that the following conditions are met:
|
|
7
|
+
|
|
8
|
+
1. Redistributions of source code must retain the above copyright notice, this
|
|
9
|
+
list of conditions and the following disclaimer.
|
|
10
|
+
|
|
11
|
+
2. Redistributions in binary form must reproduce the above copyright notice,
|
|
12
|
+
this list of conditions and the following disclaimer in the documentation
|
|
13
|
+
and/or other materials provided with the distribution.
|
|
14
|
+
|
|
15
|
+
3. Neither the name of the copyright holder nor the names of its
|
|
16
|
+
contributors may be used to endorse or promote products derived from
|
|
17
|
+
this software without specific prior written permission.
|
|
18
|
+
|
|
19
|
+
THIS SOFTWARE IS PROVIDED BY THE COPYRIGHT HOLDERS AND CONTRIBUTORS "AS IS"
|
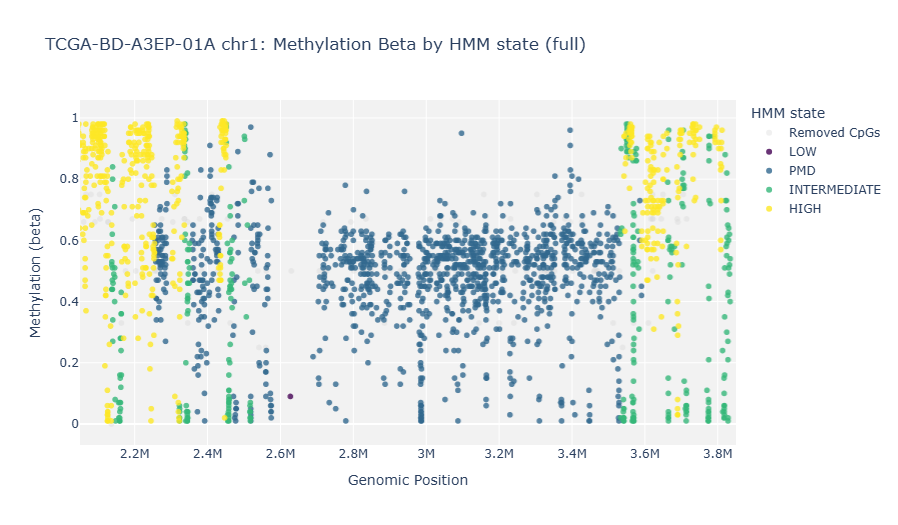
|
20
|
+
AND ANY EXPRESS OR IMPLIED WARRANTIES, INCLUDING, BUT NOT LIMITED TO, THE
|
|
21
|
+
IMPLIED WARRANTIES OF MERCHANTABILITY AND FITNESS FOR A PARTICULAR PURPOSE ARE
|
|
22
|
+
DISCLAIMED. IN NO EVENT SHALL THE COPYRIGHT HOLDER OR CONTRIBUTORS BE LIABLE
|
|
23
|
+
FOR ANY DIRECT, INDIRECT, INCIDENTAL, SPECIAL, EXEMPLARY, OR CONSEQUENTIAL
|
|
24
|
+
DAMAGES (INCLUDING, BUT NOT LIMITED TO, PROCUREMENT OF SUBSTITUTE GOODS OR
|
|
25
|
+
SERVICES; LOSS OF USE, DATA, OR PROFITS; OR BUSINESS INTERRUPTION) HOWEVER
|
|
26
|
+
CAUSED AND ON ANY THEORY OF LIABILITY, WHETHER IN CONTRACT, STRICT LIABILITY,
|
|
27
|
+
OR TORT (INCLUDING NEGLIGENCE OR OTHERWISE) ARISING IN ANY WAY OUT OF THE USE
|
|
28
|
+
OF THIS SOFTWARE, EVEN IF ADVISED OF THE POSSIBILITY OF SUCH DAMAGE.
|
methylseg-0.1.8/PKG-INFO
ADDED
|
@@ -0,0 +1,320 @@
|
|
|
1
|
+
Metadata-Version: 2.4
|
|
2
|
+
Name: methylseg
|
|
3
|
+
Version: 0.1.8
|
|
4
|
+
Summary: Methylation segmentation utilities used in the TCGA methylation analysis repo.
|
|
5
|
+
Project-URL: Documentation, https://clementlab.github.io/MethylSeg/
|
|
6
|
+
Project-URL: Repository, https://github.com/clementlab/MethylSeg
|
|
7
|
+
Requires-Python: >=3.10
|
|
8
|
+
Description-Content-Type: text/markdown
|
|
9
|
+
License-File: LICENSE
|
|
10
|
+
Requires-Dist: cthmm>=0.0.3
|
|
11
|
+
Requires-Dist: hmmlearn>=0.3.3
|
|
12
|
+
Requires-Dist: joblib>=1.5
|
|
13
|
+
Requires-Dist: matplotlib>=3.8
|
|
14
|
+
Requires-Dist: numba>=0.60
|
|
15
|
+
Requires-Dist: numpy>=1.26
|
|
16
|
+
Requires-Dist: pandas>=2.2
|
|
17
|
+
Requires-Dist: panel>=1.4
|
|
18
|
+
Requires-Dist: plotly>=5.24
|
|
19
|
+
Requires-Dist: pybedtools>=0.12.0
|
|
20
|
+
Requires-Dist: pyarrow>=17.0
|
|
21
|
+
Requires-Dist: PyYAML>=6.0
|
|
22
|
+
Requires-Dist: scikit-learn>=1.5
|
|
23
|
+
Requires-Dist: seaborn>=0.13
|
|
24
|
+
Requires-Dist: tqdm>=4.66
|
|
25
|
+
Requires-Dist: umap-learn>=0.5
|
|
26
|
+
Requires-Dist: gdown>=6.0
|
|
27
|
+
Requires-Dist: nbformat>=4.2.0
|
|
28
|
+
Provides-Extra: docs
|
|
29
|
+
Requires-Dist: ipykernel; extra == "docs"
|
|
30
|
+
Requires-Dist: myst-parser>=4.0; extra == "docs"
|
|
31
|
+
Requires-Dist: nbsphinx>=0.9.6; extra == "docs"
|
|
32
|
+
Requires-Dist: Sphinx>=7.4.7; extra == "docs"
|
|
33
|
+
Requires-Dist: sphinx-autobuild>=2024.4.16; extra == "docs"
|
|
34
|
+
Requires-Dist: sphinx-rtd-theme>=2.0.0; extra == "docs"
|
|
35
|
+
Provides-Extra: test
|
|
36
|
+
Requires-Dist: pytest>=8.0; extra == "test"
|
|
37
|
+
Dynamic: license-file
|
|
38
|
+
|
|
39
|
+
# MethylSeg
|
|
40
|
+
|
|
41
|
+
|
|
42
|
+
|
|
43
|
+
[](https://doi.org/10.5281/zenodo.22904886)
|
|
44
|
+
|
|
45
|
+
MethylSeg is a Python toolkit for identifying methylation domains from
|
|
46
|
+
whole-genome bisulfite sequencing (WGBS) and microarray methylation data. It
|
|
47
|
+
supports data preparation, methylation-state model training, genome
|
|
48
|
+
segmentation, region cleaning, and visualization.
|
|
49
|
+
|
|
50
|
+
This introduces the standard workflow. For the complete API reference
|
|
51
|
+
and example notebooks, see the [MethylSeg documentation](https://clementlab.github.io/MethylSeg/).
|
|
52
|
+
|
|
53
|
+
> [!IMPORTANT]
|
|
54
|
+
> Microarray support has currently been tested with Illumina HumanMethylation450 BeadChip (HM450K) data. Other methylation microarrays, such as Illumina HumanMethylation27 and MethylationEPIC, can be used, but array-specific defaults are not yet available and these platforms have not been formally tested.
|
|
55
|
+
|
|
56
|
+
## Hardware requirements
|
|
57
|
+
|
|
58
|
+
Memory usage depends on the input data type and dataset size. The following
|
|
59
|
+
values were observed during testing:
|
|
60
|
+
|
|
61
|
+
| Data type | Typical memory usage | Recommended system memory |
|
|
62
|
+
| --- | ---: | ---: |
|
|
63
|
+
| WGBS | ~12 GB average; up to ~15 GB peak | ≥16 GB |
|
|
64
|
+
| HM450K | <1 GB | ≥8 GB |
|
|
65
|
+
|
|
66
|
+
> [!NOTE]
|
|
67
|
+
> Numba has known compatibility issues on ARM-based systems. See the
|
|
68
|
+
> [Troubleshooting guide](https://github.com/clementlab/MethylSeg/blob/main/TROUBLESHOOTING.md)
|
|
69
|
+
> for installation guidance.
|
|
70
|
+
|
|
71
|
+
## Installation
|
|
72
|
+
|
|
73
|
+
Install from PyPI
|
|
74
|
+
|
|
75
|
+
```bash
|
|
76
|
+
python -m pip install methylseg
|
|
77
|
+
```
|
|
78
|
+
|
|
79
|
+
or install the current version from GitHub:
|
|
80
|
+
|
|
81
|
+
```bash
|
|
82
|
+
python -m pip install "git+https://github.com/clementlab/MethylSeg.git"
|
|
83
|
+
```
|
|
84
|
+
|
|
85
|
+
## Reference files
|
|
86
|
+
|
|
87
|
+
### Sample data
|
|
88
|
+
|
|
89
|
+
Three reference samples are available for use in tutorials and testing.
|
|
90
|
+
|
|
91
|
+
```bash
|
|
92
|
+
methylseg download_data_files
|
|
93
|
+
```
|
|
94
|
+
|
|
95
|
+
The two colon cancer samples were originally generated by Hansen et al. (2011) and were obtained from the data distributed with MethyLasso (Balaramane et al., 2024).
|
|
96
|
+
|
|
97
|
+
The liver cancer sample, `TCGA-BD-A3EP-01A`, was obtained from the [TCGA-LIHC project through the NCI Genomic Data Commons](https://portal.gdc.cancer.gov/cases/de302b98-250d-42fe-9945-11e1b2bbd3c6).
|
|
98
|
+
|
|
99
|
+
| Sample name | WGBS filename | HM450K filename |
|
|
100
|
+
| --- | --- | --- |
|
|
101
|
+
| `WGBS_colon-primary-tumor_1` | `WGBS_colon-primary-tumor_1_wgbs.tsv` | `WGBS_colon-primary-tumor_1_450k.beta` |
|
|
102
|
+
| `WGBS_colon-primary-tumor_2` | `WGBS_colon-primary-tumor_2_wgbs.tsv` | `WGBS_colon-primary-tumor_2_450k.beta` |
|
|
103
|
+
| `TCGA-BD-A3EP-01A` | N/A | `TCGA-BD-A3EP-01A_450k.tsv` |
|
|
104
|
+
|
|
105
|
+
#### References
|
|
106
|
+
|
|
107
|
+
1. Hansen, K. D., et al. Increased methylation variation in epigenetic domains across cancer types. *Nature Genetics* **43**, 768–775 (2011).
|
|
108
|
+
2. Balaramane, D., Spill, Y. G., Weber, M. & Bardet, A. F. MethyLasso: a segmentation approach to analyze DNA methylation patterns and identify differentially methylated regions from whole-genome datasets. *Nucleic Acids Research* **52**, e98 (2024).
|
|
109
|
+
|
|
110
|
+
### Pretrained models
|
|
111
|
+
|
|
112
|
+
Two pretrained models are provided for tutorials, testing, and exploratory
|
|
113
|
+
analysis:
|
|
114
|
+
|
|
115
|
+
| Model filename | Training data |
|
|
116
|
+
| --- | --- |
|
|
117
|
+
| `wgbs_colon_model` | `WGBS_colon-primary-tumor_1_wgbs.tsv` |
|
|
118
|
+
| `tcga_hm450k_model` | `TCGA-BD-A3EP-01A_450k.tsv` |
|
|
119
|
+
|
|
120
|
+
Load a pretrained model with `get_pretrained_model`:
|
|
121
|
+
|
|
122
|
+
```python
|
|
123
|
+
# WGBS model
|
|
124
|
+
wgbs_saved = MethylSegPathway.get_pretrained_model(
|
|
125
|
+
out_dir=OUTPUT_DIR,
|
|
126
|
+
resolution="wgbs",
|
|
127
|
+
)
|
|
128
|
+
|
|
129
|
+
# HM450K model
|
|
130
|
+
hm450k_saved = MethylSegPathway.get_pretrained_model(
|
|
131
|
+
out_dir=OUTPUT_DIR,
|
|
132
|
+
resolution="450k",
|
|
133
|
+
)
|
|
134
|
+
```
|
|
135
|
+
|
|
136
|
+
## Quickstart
|
|
137
|
+
|
|
138
|
+
The default workflow trains a model on your input, segments the sample,
|
|
139
|
+
cleans the calls, and draws a cleaned methylation state overlay.
|
|
140
|
+
|
|
141
|
+
Use `resolution="wgbs"` for WGBS count tables or `resolution="450k"` for HM450K
|
|
142
|
+
beta-value tables. Other microarray platforms may require manually configured
|
|
143
|
+
parameters until array-specific defaults are added.
|
|
144
|
+
|
|
145
|
+
```python
|
|
146
|
+
from pathlib import Path
|
|
147
|
+
|
|
148
|
+
from methylseg import MethylSegPathway, MethylationStates
|
|
149
|
+
from methylseg.helper_classes import DATA_DIR
|
|
150
|
+
|
|
151
|
+
reference_dir = DATA_DIR / "reference_files"
|
|
152
|
+
|
|
153
|
+
# WGBS: replace these with your own sample name and count table.
|
|
154
|
+
# sample_name = "WGBS_colon-primary-tumor_1"
|
|
155
|
+
# sample_file = reference_dir / "WGBS_colon-primary-tumor_1_wgbs.tsv.gz"
|
|
156
|
+
# resolution = "wgbs"
|
|
157
|
+
|
|
158
|
+
# TCGA/HM450K alternative:
|
|
159
|
+
sample_name = "TCGA-BD-A3EP-01A"
|
|
160
|
+
sample_file = reference_dir / "TCGA-BD-A3EP-01A_450k.tsv.gz"
|
|
161
|
+
resolution = "450k"
|
|
162
|
+
|
|
163
|
+
sample_info, removed_df = MethylSegPathway.prepare_sample_info(
|
|
164
|
+
sample_name=sample_name,
|
|
165
|
+
sample_file=sample_file,
|
|
166
|
+
resolution=resolution,
|
|
167
|
+
remove_low_coverage_like_cpgs=True,
|
|
168
|
+
)
|
|
169
|
+
|
|
170
|
+
pathway = MethylSegPathway(
|
|
171
|
+
train_sample_info=sample_info,
|
|
172
|
+
out_dir=Path("out/quickstart") / sample_name,
|
|
173
|
+
)
|
|
174
|
+
|
|
175
|
+
pathway.run_pathway()
|
|
176
|
+
|
|
177
|
+
fig = pathway.plot_labels(
|
|
178
|
+
sample_info=sample_info,
|
|
179
|
+
sample_info_removed=removed_df,
|
|
180
|
+
chrom="chr1",
|
|
181
|
+
region_start=2_200_000,
|
|
182
|
+
region_end=3_700_000
|
|
183
|
+
)
|
|
184
|
+
```
|
|
185
|
+
|
|
186
|
+
|
|
187
|
+
|
|
188
|
+
`run_pathway(sample_info=sample_info, chroms=["chr1"])` performs fitting,
|
|
189
|
+
segmentation, cleaning, and summary-file writing in one call.
|
|
190
|
+
|
|
191
|
+
## Input formats
|
|
192
|
+
|
|
193
|
+
MethylSeg accepts tab-delimited `.tsv` and `.tsv.gz` files containing DNA methylation measurements. Genomic coordinates must use the 0-based, half-open coordinate system, and chromosome names must be consistent throughout each file.
|
|
194
|
+
|
|
195
|
+
Two BED-like input formats are supported.
|
|
196
|
+
|
|
197
|
+
### Beta-value format
|
|
198
|
+
|
|
199
|
+
This format can be used for either WGBS or microarray data. It contains four required columns:
|
|
200
|
+
|
|
201
|
+
1. chromosome;
|
|
202
|
+
2. CpG start position;
|
|
203
|
+
3. CpG end position; and
|
|
204
|
+
4. methylation beta value.
|
|
205
|
+
|
|
206
|
+
For microarray data, an optional fifth column containing the probe identifier may be included. Probe identifiers are retained as metadata.
|
|
207
|
+
|
|
208
|
+
| CpG_chrm | CpG_beg | CpG_end | beta | probe |
|
|
209
|
+
| -------- | ------: | ------: | ---: | ---------- |
|
|
210
|
+
| chr1 | 15864 | 15866 | 0.0 | cg13869341 |
|
|
211
|
+
| chr1 | 29406 | 29408 | 0.0 | cg12045430 |
|
|
212
|
+
| chr1 | 29424 | 29426 | 0.0 | cg20826792 |
|
|
213
|
+
|
|
214
|
+
|
|
215
|
+
### WGBS count format
|
|
216
|
+
|
|
217
|
+
This format contains five required columns:
|
|
218
|
+
|
|
219
|
+
1. chromosome;
|
|
220
|
+
2. CpG start position;
|
|
221
|
+
3. CpG end position;
|
|
222
|
+
4. methylated read count; and
|
|
223
|
+
5. total read coverage.
|
|
224
|
+
|
|
225
|
+
MethylSeg calculates the beta value internally as:
|
|
226
|
+
$
|
|
227
|
+
\mathrm{beta} = \frac{\mathrm{methylated\ reads}}{\mathrm{total\ coverage}}.
|
|
228
|
+
$
|
|
229
|
+
|
|
230
|
+
| CpG_chrm | CpG_beg | CpG_end | meth | coverage |
|
|
231
|
+
| -------- | ------: | ------: | ---: | -------: |
|
|
232
|
+
| chr1 | 10468 | 10470 | 14 | 15 |
|
|
233
|
+
| chr1 | 10470 | 10472 | 10 | 10 |
|
|
234
|
+
| chr1 | 10483 | 10485 | 23 | 28 |
|
|
235
|
+
|
|
236
|
+
|
|
237
|
+
## Outputs
|
|
238
|
+
|
|
239
|
+
The final output of MethylSeg consists primarily of BED files containing the
|
|
240
|
+
identified methylation domains. Separate files are generated for each
|
|
241
|
+
methylation state: `HIGH`, `INTERMEDIATE`, `LOW`, and `PMD`.
|
|
242
|
+
|
|
243
|
+
For each chromosome and state, MethylSeg writes:
|
|
244
|
+
|
|
245
|
+
| Output | Filename pattern | Description |
|
|
246
|
+
| --- | --- | --- |
|
|
247
|
+
| Raw regions | `segments_{chrom}_{sample_id}_{resolution}_{state}.bed` | Regions produced directly by genome segmentation |
|
|
248
|
+
| Cleaned regions | `clean_regions/segments_cleaned_{chrom}_{sample_id}_{resolution}_{state}.bed` | Regions retained after merging and filtering |
|
|
249
|
+
| Cleaned metadata | `clean_regions/metadata_cleaned_{chrom}_{sample_id}_{resolution}_{state}.tsv` | Additional information about the cleaned regions |
|
|
250
|
+
|
|
251
|
+
After all requested chromosomes have been processed, MethylSeg also creates
|
|
252
|
+
genome-wide files for each state under `summary_files/`:
|
|
253
|
+
|
|
254
|
+
| Output | Filename pattern |
|
|
255
|
+
| --- | --- |
|
|
256
|
+
| Genome-wide raw regions | `segments_raw_{state}.bed` |
|
|
257
|
+
| Genome-wide cleaned regions | `segments_cleaned_{state}.bed` |
|
|
258
|
+
| Genome-wide cleaned metadata | `metadata_cleaned_{state}.tsv` |
|
|
259
|
+
|
|
260
|
+
A simplified output directory has the following structure:
|
|
261
|
+
|
|
262
|
+
```text
|
|
263
|
+
methylseg_output/
|
|
264
|
+
├── segments_chr1_sample.wgbs_PMD.bed
|
|
265
|
+
├── segments_chr1_sample.wgbs_LOW.bed
|
|
266
|
+
├── clean_regions/
|
|
267
|
+
│ ├── segments_cleaned_chr1_sample.wgbs_PMD.bed
|
|
268
|
+
│ ├── segments_cleaned_chr1_sample.wgbs_LOW.bed
|
|
269
|
+
│ ├── metadata_cleaned_chr1_sample.wgbs_PMD.tsv
|
|
270
|
+
│ └── metadata_cleaned_chr1_sample.wgbs_LOW.tsv
|
|
271
|
+
└── summary_files/
|
|
272
|
+
├── segments_raw_PMD.bed
|
|
273
|
+
├── segments_raw_LOW.bed
|
|
274
|
+
├── segments_cleaned_PMD.bed
|
|
275
|
+
├── segments_cleaned_LOW.bed
|
|
276
|
+
├── metadata_cleaned_PMD.tsv
|
|
277
|
+
└── metadata_cleaned_LOW.tsv
|
|
278
|
+
```
|
|
279
|
+
|
|
280
|
+
Each BED file contains 0-based, half-open genomic coordinates and the assigned
|
|
281
|
+
methylation state. For example, segments_cleaned_PMD.bed may contain:
|
|
282
|
+
|
|
283
|
+
| Chromosome | Start | End | State |
|
|
284
|
+
| ---------- | --------: | --------: | ----- |
|
|
285
|
+
| `chr1` | 1,261,344 | 1,323,691 | `PMD` |
|
|
286
|
+
| `chr1` | 2,789,167 | 2,999,312 | `PMD` |
|
|
287
|
+
| `chr1` | 3,885,138 | 3,916,529 | `PMD` |
|
|
288
|
+
| `chr1` | 6,167,866 | 6,186,177 | `PMD` |
|
|
289
|
+
|
|
290
|
+
The BED files themselves are tab-delimited and do not contain a header.
|
|
291
|
+
|
|
292
|
+
## Citation
|
|
293
|
+
|
|
294
|
+
If you use MethylSeg, please cite the software using the
|
|
295
|
+
[CITATION.cff](https://github.com/clementlab/MethylSeg/blob/main/CITATION.cff)
|
|
296
|
+
metadata. On GitHub, select **Cite this repository** to copy the citation in APA
|
|
297
|
+
or BibTeX format.
|
|
298
|
+
|
|
299
|
+
A manuscript describing MethylSeg is in preparation. Its citation will be added
|
|
300
|
+
when available.
|
|
301
|
+
|
|
302
|
+
## Planned support
|
|
303
|
+
|
|
304
|
+
See [ROADMAP.md](https://github.com/clementlab/MethylSeg/blob/main/ROADMAP.md)
|
|
305
|
+
|
|
306
|
+
## Reporting issues
|
|
307
|
+
|
|
308
|
+
To report a bug, request a feature, or ask a question about MethylSeg, open an
|
|
309
|
+
issue on the [GitHub issue tracker](https://github.com/clementlab/MethylSeg/issues).
|
|
310
|
+
|
|
311
|
+
When reporting a bug, please include:
|
|
312
|
+
|
|
313
|
+
- Your MethylSeg version
|
|
314
|
+
- Your Python version and operating system
|
|
315
|
+
- The complete error message or traceback
|
|
316
|
+
- A minimal example that reproduces the problem, when possible
|
|
317
|
+
|
|
318
|
+
## License
|
|
319
|
+
|
|
320
|
+
MethylSeg is distributed under the [BSD 3-Clause License](LICENSE.md).
|
|
@@ -0,0 +1,282 @@
|
|
|
1
|
+
# MethylSeg
|
|
2
|
+
|
|
3
|
+

|
|
4
|
+
|
|
5
|
+
[](https://doi.org/10.5281/zenodo.22904886)
|
|
6
|
+
|
|
7
|
+
MethylSeg is a Python toolkit for identifying methylation domains from
|
|
8
|
+
whole-genome bisulfite sequencing (WGBS) and microarray methylation data. It
|
|
9
|
+
supports data preparation, methylation-state model training, genome
|
|
10
|
+
segmentation, region cleaning, and visualization.
|
|
11
|
+
|
|
12
|
+
This introduces the standard workflow. For the complete API reference
|
|
13
|
+
and example notebooks, see the [MethylSeg documentation](https://clementlab.github.io/MethylSeg/).
|
|
14
|
+
|
|
15
|
+
> [!IMPORTANT]
|
|
16
|
+
> Microarray support has currently been tested with Illumina HumanMethylation450 BeadChip (HM450K) data. Other methylation microarrays, such as Illumina HumanMethylation27 and MethylationEPIC, can be used, but array-specific defaults are not yet available and these platforms have not been formally tested.
|
|
17
|
+
|
|
18
|
+
## Hardware requirements
|
|
19
|
+
|
|
20
|
+
Memory usage depends on the input data type and dataset size. The following
|
|
21
|
+
values were observed during testing:
|
|
22
|
+
|
|
23
|
+
| Data type | Typical memory usage | Recommended system memory |
|
|
24
|
+
| --- | ---: | ---: |
|
|
25
|
+
| WGBS | ~12 GB average; up to ~15 GB peak | ≥16 GB |
|
|
26
|
+
| HM450K | <1 GB | ≥8 GB |
|
|
27
|
+
|
|
28
|
+
> [!NOTE]
|
|
29
|
+
> Numba has known compatibility issues on ARM-based systems. See the
|
|
30
|
+
> [Troubleshooting guide](https://github.com/clementlab/MethylSeg/blob/main/TROUBLESHOOTING.md)
|
|
31
|
+
> for installation guidance.
|
|
32
|
+
|
|
33
|
+
## Installation
|
|
34
|
+
|
|
35
|
+
Install from PyPI
|
|
36
|
+
|
|
37
|
+
```bash
|
|
38
|
+
python -m pip install methylseg
|
|
39
|
+
```
|
|
40
|
+
|
|
41
|
+
or install the current version from GitHub:
|
|
42
|
+
|
|
43
|
+
```bash
|
|
44
|
+
python -m pip install "git+https://github.com/clementlab/MethylSeg.git"
|
|
45
|
+
```
|
|
46
|
+
|
|
47
|
+
## Reference files
|
|
48
|
+
|
|
49
|
+
### Sample data
|
|
50
|
+
|
|
51
|
+
Three reference samples are available for use in tutorials and testing.
|
|
52
|
+
|
|
53
|
+
```bash
|
|
54
|
+
methylseg download_data_files
|
|
55
|
+
```
|
|
56
|
+
|
|
57
|
+
The two colon cancer samples were originally generated by Hansen et al. (2011) and were obtained from the data distributed with MethyLasso (Balaramane et al., 2024).
|
|
58
|
+
|
|
59
|
+
The liver cancer sample, `TCGA-BD-A3EP-01A`, was obtained from the [TCGA-LIHC project through the NCI Genomic Data Commons](https://portal.gdc.cancer.gov/cases/de302b98-250d-42fe-9945-11e1b2bbd3c6).
|
|
60
|
+
|
|
61
|
+
| Sample name | WGBS filename | HM450K filename |
|
|
62
|
+
| --- | --- | --- |
|
|
63
|
+
| `WGBS_colon-primary-tumor_1` | `WGBS_colon-primary-tumor_1_wgbs.tsv` | `WGBS_colon-primary-tumor_1_450k.beta` |
|
|
64
|
+
| `WGBS_colon-primary-tumor_2` | `WGBS_colon-primary-tumor_2_wgbs.tsv` | `WGBS_colon-primary-tumor_2_450k.beta` |
|
|
65
|
+
| `TCGA-BD-A3EP-01A` | N/A | `TCGA-BD-A3EP-01A_450k.tsv` |
|
|
66
|
+
|
|
67
|
+
#### References
|
|
68
|
+
|
|
69
|
+
1. Hansen, K. D., et al. Increased methylation variation in epigenetic domains across cancer types. *Nature Genetics* **43**, 768–775 (2011).
|
|
70
|
+
2. Balaramane, D., Spill, Y. G., Weber, M. & Bardet, A. F. MethyLasso: a segmentation approach to analyze DNA methylation patterns and identify differentially methylated regions from whole-genome datasets. *Nucleic Acids Research* **52**, e98 (2024).
|
|
71
|
+
|
|
72
|
+
### Pretrained models
|
|
73
|
+
|
|
74
|
+
Two pretrained models are provided for tutorials, testing, and exploratory
|
|
75
|
+
analysis:
|
|
76
|
+
|
|
77
|
+
| Model filename | Training data |
|
|
78
|
+
| --- | --- |
|
|
79
|
+
| `wgbs_colon_model` | `WGBS_colon-primary-tumor_1_wgbs.tsv` |
|
|
80
|
+
| `tcga_hm450k_model` | `TCGA-BD-A3EP-01A_450k.tsv` |
|
|
81
|
+
|
|
82
|
+
Load a pretrained model with `get_pretrained_model`:
|
|
83
|
+
|
|
84
|
+
```python
|
|
85
|
+
# WGBS model
|
|
86
|
+
wgbs_saved = MethylSegPathway.get_pretrained_model(
|
|
87
|
+
out_dir=OUTPUT_DIR,
|
|
88
|
+
resolution="wgbs",
|
|
89
|
+
)
|
|
90
|
+
|
|
91
|
+
# HM450K model
|
|
92
|
+
hm450k_saved = MethylSegPathway.get_pretrained_model(
|
|
93
|
+
out_dir=OUTPUT_DIR,
|
|
94
|
+
resolution="450k",
|
|
95
|
+
)
|
|
96
|
+
```
|
|
97
|
+
|
|
98
|
+
## Quickstart
|
|
99
|
+
|
|
100
|
+
The default workflow trains a model on your input, segments the sample,
|
|
101
|
+
cleans the calls, and draws a cleaned methylation state overlay.
|
|
102
|
+
|
|
103
|
+
Use `resolution="wgbs"` for WGBS count tables or `resolution="450k"` for HM450K
|
|
104
|
+
beta-value tables. Other microarray platforms may require manually configured
|
|
105
|
+
parameters until array-specific defaults are added.
|
|
106
|
+
|
|
107
|
+
```python
|
|
108
|
+
from pathlib import Path
|
|
109
|
+
|
|
110
|
+
from methylseg import MethylSegPathway, MethylationStates
|
|
111
|
+
from methylseg.helper_classes import DATA_DIR
|
|
112
|
+
|
|
113
|
+
reference_dir = DATA_DIR / "reference_files"
|
|
114
|
+
|
|
115
|
+
# WGBS: replace these with your own sample name and count table.
|
|
116
|
+
# sample_name = "WGBS_colon-primary-tumor_1"
|
|
117
|
+
# sample_file = reference_dir / "WGBS_colon-primary-tumor_1_wgbs.tsv.gz"
|
|
118
|
+
# resolution = "wgbs"
|
|
119
|
+
|
|
120
|
+
# TCGA/HM450K alternative:
|
|
121
|
+
sample_name = "TCGA-BD-A3EP-01A"
|
|
122
|
+
sample_file = reference_dir / "TCGA-BD-A3EP-01A_450k.tsv.gz"
|
|
123
|
+
resolution = "450k"
|
|
124
|
+
|
|
125
|
+
sample_info, removed_df = MethylSegPathway.prepare_sample_info(
|
|
126
|
+
sample_name=sample_name,
|
|
127
|
+
sample_file=sample_file,
|
|
128
|
+
resolution=resolution,
|
|
129
|
+
remove_low_coverage_like_cpgs=True,
|
|
130
|
+
)
|
|
131
|
+
|
|
132
|
+
pathway = MethylSegPathway(
|
|
133
|
+
train_sample_info=sample_info,
|
|
134
|
+
out_dir=Path("out/quickstart") / sample_name,
|
|
135
|
+
)
|
|
136
|
+
|
|
137
|
+
pathway.run_pathway()
|
|
138
|
+
|
|
139
|
+
fig = pathway.plot_labels(
|
|
140
|
+
sample_info=sample_info,
|
|
141
|
+
sample_info_removed=removed_df,
|
|
142
|
+
chrom="chr1",
|
|
143
|
+
region_start=2_200_000,
|
|
144
|
+
region_end=3_700_000
|
|
145
|
+
)
|
|
146
|
+
```
|
|
147
|
+
|
|
148
|
+

|
|
149
|
+
|
|
150
|
+
`run_pathway(sample_info=sample_info, chroms=["chr1"])` performs fitting,
|
|
151
|
+
segmentation, cleaning, and summary-file writing in one call.
|
|
152
|
+
|
|
153
|
+
## Input formats
|
|
154
|
+
|
|
155
|
+
MethylSeg accepts tab-delimited `.tsv` and `.tsv.gz` files containing DNA methylation measurements. Genomic coordinates must use the 0-based, half-open coordinate system, and chromosome names must be consistent throughout each file.
|
|
156
|
+
|
|
157
|
+
Two BED-like input formats are supported.
|
|
158
|
+
|
|
159
|
+
### Beta-value format
|
|
160
|
+
|
|
161
|
+
This format can be used for either WGBS or microarray data. It contains four required columns:
|
|
162
|
+
|
|
163
|
+
1. chromosome;
|
|
164
|
+
2. CpG start position;
|
|
165
|
+
3. CpG end position; and
|
|
166
|
+
4. methylation beta value.
|
|
167
|
+
|
|
168
|
+
For microarray data, an optional fifth column containing the probe identifier may be included. Probe identifiers are retained as metadata.
|
|
169
|
+
|
|
170
|
+
| CpG_chrm | CpG_beg | CpG_end | beta | probe |
|
|
171
|
+
| -------- | ------: | ------: | ---: | ---------- |
|
|
172
|
+
| chr1 | 15864 | 15866 | 0.0 | cg13869341 |
|
|
173
|
+
| chr1 | 29406 | 29408 | 0.0 | cg12045430 |
|
|
174
|
+
| chr1 | 29424 | 29426 | 0.0 | cg20826792 |
|
|
175
|
+
|
|
176
|
+
|
|
177
|
+
### WGBS count format
|
|
178
|
+
|
|
179
|
+
This format contains five required columns:
|
|
180
|
+
|
|
181
|
+
1. chromosome;
|
|
182
|
+
2. CpG start position;
|
|
183
|
+
3. CpG end position;
|
|
184
|
+
4. methylated read count; and
|
|
185
|
+
5. total read coverage.
|
|
186
|
+
|
|
187
|
+
MethylSeg calculates the beta value internally as:
|
|
188
|
+
$
|
|
189
|
+
\mathrm{beta} = \frac{\mathrm{methylated\ reads}}{\mathrm{total\ coverage}}.
|
|
190
|
+
$
|
|
191
|
+
|
|
192
|
+
| CpG_chrm | CpG_beg | CpG_end | meth | coverage |
|
|
193
|
+
| -------- | ------: | ------: | ---: | -------: |
|
|
194
|
+
| chr1 | 10468 | 10470 | 14 | 15 |
|
|
195
|
+
| chr1 | 10470 | 10472 | 10 | 10 |
|
|
196
|
+
| chr1 | 10483 | 10485 | 23 | 28 |
|
|
197
|
+
|
|
198
|
+
|
|
199
|
+
## Outputs
|
|
200
|
+
|
|
201
|
+
The final output of MethylSeg consists primarily of BED files containing the
|
|
202
|
+
identified methylation domains. Separate files are generated for each
|
|
203
|
+
methylation state: `HIGH`, `INTERMEDIATE`, `LOW`, and `PMD`.
|
|
204
|
+
|
|
205
|
+
For each chromosome and state, MethylSeg writes:
|
|
206
|
+
|
|
207
|
+
| Output | Filename pattern | Description |
|
|
208
|
+
| --- | --- | --- |
|
|
209
|
+
| Raw regions | `segments_{chrom}_{sample_id}_{resolution}_{state}.bed` | Regions produced directly by genome segmentation |
|
|
210
|
+
| Cleaned regions | `clean_regions/segments_cleaned_{chrom}_{sample_id}_{resolution}_{state}.bed` | Regions retained after merging and filtering |
|
|
211
|
+
| Cleaned metadata | `clean_regions/metadata_cleaned_{chrom}_{sample_id}_{resolution}_{state}.tsv` | Additional information about the cleaned regions |
|
|
212
|
+
|
|
213
|
+
After all requested chromosomes have been processed, MethylSeg also creates
|
|
214
|
+
genome-wide files for each state under `summary_files/`:
|
|
215
|
+
|
|
216
|
+
| Output | Filename pattern |
|
|
217
|
+
| --- | --- |
|
|
218
|
+
| Genome-wide raw regions | `segments_raw_{state}.bed` |
|
|
219
|
+
| Genome-wide cleaned regions | `segments_cleaned_{state}.bed` |
|
|
220
|
+
| Genome-wide cleaned metadata | `metadata_cleaned_{state}.tsv` |
|
|
221
|
+
|
|
222
|
+
A simplified output directory has the following structure:
|
|
223
|
+
|
|
224
|
+
```text
|
|
225
|
+
methylseg_output/
|
|
226
|
+
├── segments_chr1_sample.wgbs_PMD.bed
|
|
227
|
+
├── segments_chr1_sample.wgbs_LOW.bed
|
|
228
|
+
├── clean_regions/
|
|
229
|
+
│ ├── segments_cleaned_chr1_sample.wgbs_PMD.bed
|
|
230
|
+
│ ├── segments_cleaned_chr1_sample.wgbs_LOW.bed
|
|
231
|
+
│ ├── metadata_cleaned_chr1_sample.wgbs_PMD.tsv
|
|
232
|
+
│ └── metadata_cleaned_chr1_sample.wgbs_LOW.tsv
|
|
233
|
+
└── summary_files/
|
|
234
|
+
├── segments_raw_PMD.bed
|
|
235
|
+
├── segments_raw_LOW.bed
|
|
236
|
+
├── segments_cleaned_PMD.bed
|
|
237
|
+
├── segments_cleaned_LOW.bed
|
|
238
|
+
├── metadata_cleaned_PMD.tsv
|
|
239
|
+
└── metadata_cleaned_LOW.tsv
|
|
240
|
+
```
|
|
241
|
+
|
|
242
|
+
Each BED file contains 0-based, half-open genomic coordinates and the assigned
|
|
243
|
+
methylation state. For example, segments_cleaned_PMD.bed may contain:
|
|
244
|
+
|
|
245
|
+
| Chromosome | Start | End | State |
|
|
246
|
+
| ---------- | --------: | --------: | ----- |
|
|
247
|
+
| `chr1` | 1,261,344 | 1,323,691 | `PMD` |
|
|
248
|
+
| `chr1` | 2,789,167 | 2,999,312 | `PMD` |
|
|
249
|
+
| `chr1` | 3,885,138 | 3,916,529 | `PMD` |
|
|
250
|
+
| `chr1` | 6,167,866 | 6,186,177 | `PMD` |
|
|
251
|
+
|
|
252
|
+
The BED files themselves are tab-delimited and do not contain a header.
|
|
253
|
+
|
|
254
|
+
## Citation
|
|
255
|
+
|
|
256
|
+
If you use MethylSeg, please cite the software using the
|
|
257
|
+
[CITATION.cff](https://github.com/clementlab/MethylSeg/blob/main/CITATION.cff)
|
|
258
|
+
metadata. On GitHub, select **Cite this repository** to copy the citation in APA
|
|
259
|
+
or BibTeX format.
|
|
260
|
+
|
|
261
|
+
A manuscript describing MethylSeg is in preparation. Its citation will be added
|
|
262
|
+
when available.
|
|
263
|
+
|
|
264
|
+
## Planned support
|
|
265
|
+
|
|
266
|
+
See [ROADMAP.md](https://github.com/clementlab/MethylSeg/blob/main/ROADMAP.md)
|
|
267
|
+
|
|
268
|
+
## Reporting issues
|
|
269
|
+
|
|
270
|
+
To report a bug, request a feature, or ask a question about MethylSeg, open an
|
|
271
|
+
issue on the [GitHub issue tracker](https://github.com/clementlab/MethylSeg/issues).
|
|
272
|
+
|
|
273
|
+
When reporting a bug, please include:
|
|
274
|
+
|
|
275
|
+
- Your MethylSeg version
|
|
276
|
+
- Your Python version and operating system
|
|
277
|
+
- The complete error message or traceback
|
|
278
|
+
- A minimal example that reproduces the problem, when possible
|
|
279
|
+
|
|
280
|
+
## License
|
|
281
|
+
|
|
282
|
+
MethylSeg is distributed under the [BSD 3-Clause License](LICENSE.md).
|
|
@@ -0,0 +1,38 @@
|
|
|
1
|
+
"""Public package exports for the reusable methylseg workflow."""
|
|
2
|
+
|
|
3
|
+
from .helper_classes import (
|
|
4
|
+
MethylStateAssignmentMethod,
|
|
5
|
+
MethylationStates,
|
|
6
|
+
MethylDataPrep,
|
|
7
|
+
SampleInfo,
|
|
8
|
+
HMMType,
|
|
9
|
+
)
|
|
10
|
+
from .methylseg_hmm import (
|
|
11
|
+
CTMethylSegHMM,
|
|
12
|
+
MethylSegHMM,
|
|
13
|
+
StickyCategoricalMethylSegHMM,
|
|
14
|
+
)
|
|
15
|
+
from .methyl_state_assigner import MethylStateAssigner
|
|
16
|
+
from .methyl_state_analyzer import MethylStateAnalyzer
|
|
17
|
+
from .methyl_segmentor import MethylSegmentor
|
|
18
|
+
from .methylseg_pathway import MethylSegPathway
|
|
19
|
+
from .methylseg_config import MethylSegConfig
|
|
20
|
+
from .utils import get_biological_state_colors, get_cluster_colors
|
|
21
|
+
|
|
22
|
+
__all__ = [
|
|
23
|
+
"CTMethylSegHMM",
|
|
24
|
+
"MethylDataPrep",
|
|
25
|
+
"MethylSegConfig",
|
|
26
|
+
"MethylSegHMM",
|
|
27
|
+
"MethylSegPathway",
|
|
28
|
+
"MethylSegmentor",
|
|
29
|
+
"MethylStateAnalyzer",
|
|
30
|
+
"MethylStateAssigner",
|
|
31
|
+
"MethylStateAssignmentMethod",
|
|
32
|
+
"MethylationStates",
|
|
33
|
+
"HMMType",
|
|
34
|
+
"SampleInfo",
|
|
35
|
+
"StickyCategoricalMethylSegHMM",
|
|
36
|
+
"get_biological_state_colors",
|
|
37
|
+
"get_cluster_colors",
|
|
38
|
+
]
|
|
@@ -0,0 +1,32 @@
|
|
|
1
|
+
from .data_manager import download_data_files
|
|
2
|
+
import argparse
|
|
3
|
+
|
|
4
|
+
#TODO - add subcommands for pathway running, segmentation and plotting
|
|
5
|
+
def main():
|
|
6
|
+
"""
|
|
7
|
+
Run the ``methylseg`` command-line interface.
|
|
8
|
+
|
|
9
|
+
Returns
|
|
10
|
+
-------
|
|
11
|
+
None
|
|
12
|
+
Parses CLI arguments, executes the requested subcommand, and prints
|
|
13
|
+
help text when no subcommand is provided.
|
|
14
|
+
"""
|
|
15
|
+
parser = argparse.ArgumentParser(prog="methylseg")
|
|
16
|
+
|
|
17
|
+
subparsers = parser.add_subparsers(dest="command")
|
|
18
|
+
|
|
19
|
+
# subcommand: download_data_files
|
|
20
|
+
dl_parser = subparsers.add_parser("download_data_files")
|
|
21
|
+
dl_parser.add_argument(
|
|
22
|
+
"--cleanup_existing",
|
|
23
|
+
action="store_true",
|
|
24
|
+
help="Delete existing files before downloading",
|
|
25
|
+
)
|
|
26
|
+
|
|
27
|
+
args = parser.parse_args()
|
|
28
|
+
|
|
29
|
+
if args.command == "download_data_files":
|
|
30
|
+
download_data_files(cleanup_existing=args.cleanup_existing)
|
|
31
|
+
else:
|
|
32
|
+
parser.print_help()
|